Loading...

Current Issue

    Chinese Journal of Catalysis
    2027, Vol. 92
    Online: 18 January 2027

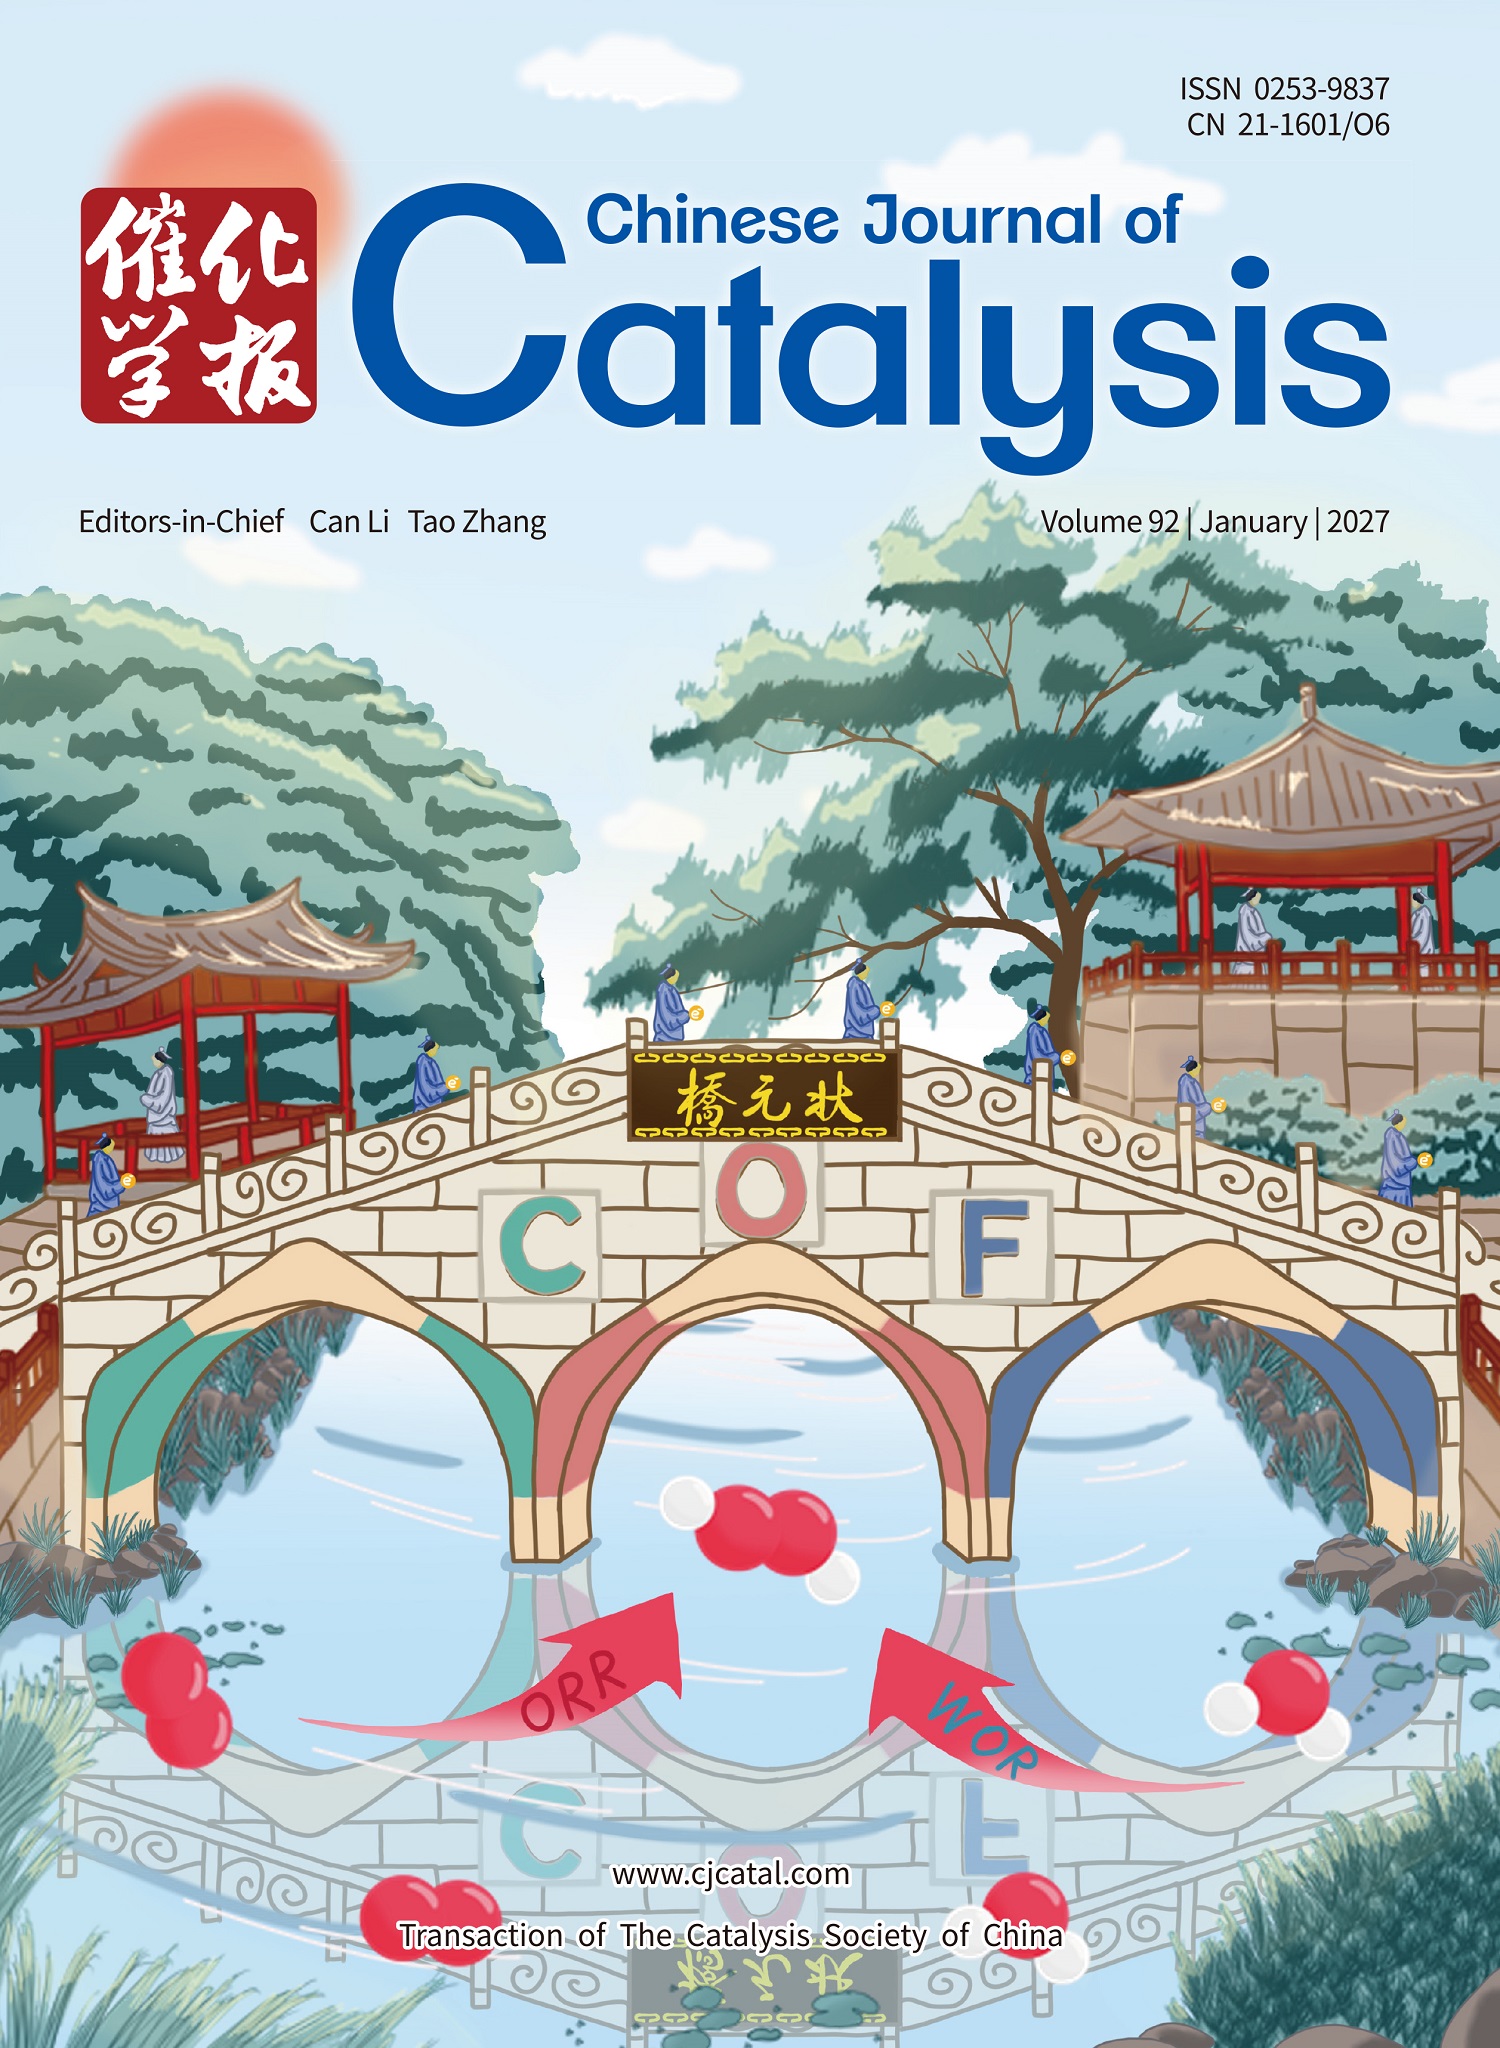
    Cover: Profs. Zhonghua Li, Feng-Ming Zhang, Yang Liu, Qichun Zhang, and coworkers in their article on pages 130–144 reported a series of isostructural covalent organic frameworks (COFs) for solar-driven H2O2 photosynthesis. By coupling global D3h framework symmetry with local electronic asymmetry, the triazine-based TAT-COF generates an internal electric field for ultrafast exciton dissociation while maintaining ordered long-range charge transport, achieving high photocatalytic activity through a dual-pathway mechanism. This work provides an effective molecular design strategy for artificial photosynthesis.
    For Selected: Toggle Thumbnails
    Reviews
    S-scheme heterojunctions for coupled redox photocatalysis: Orchestrating electron-hole synergy to unlock dual half-reactions
    Banghu Wei, Wei Ren, Jinhe Li, Weikang Wang, Qinqin Liu
    2027, 92:  1-36.  DOI: 10.1016/S1872-2067(26)65276-X
    Abstract ( 35 )   HTML ( 2 )   PDF (29819KB) ( 8 )  

    Against the backdrop of the global "dual-carbon" strategy, solar-driven photocatalysis emerges as a promising avenue for clean energy conversion and carbon neutralization. However, conventional single photocatalysts suffer from rapid carrier recombination and mismatched redox reaction kinetics, severely limiting their catalytic performance. Different from conventional heterojunctions that compromise redox potential for enhanced charge separation, S-scheme heterojunctions feature a unique band alignment that selectively eliminates useless low-energy carriers via interfacial recombination. This configuration preserves high-energy electrons and holes with robust redox capabilities, fundamentally resolving the intrinsic trade-off between charge separation efficiency and thermodynamic redox driving force in photocatalysis. Based on this working principle, this review systematically elucidates the efficient utilization mechanism of electron-hole pairs in S-scheme heterojunctions, with a specific focus on the synergistic promotion mechanism between paired reduction and oxidation half-reactions in photocatalytic redox coupling systems. We highlight three major categories of reduction reactions—hydrogen production, CO2 reduction, and H2O2—coupled with diversified value-added oxidation processes (e.g., organic molecule oxidation, biomass upgrading, and pollutant oxidative degradation). Through the rational regulation of active site construction, band structure engineering, and interfacial charge modulation in S-scheme heterojunctions, the reaction kinetics, conversion efficiency, and catalytic pathways of both reduction and oxidation half-reactions can be synchronously optimized, which substantially improves the atom utilization efficiency and overall economic value of solar energy photocatalytic conversion. This review systematically clarifies the intrinsic correlation and synergistic mechanism between S-scheme heterojunction design and dual-reaction coupling catalysis, aiming to provide feasible theoretical guidance and technical roadmaps for the rational construction of high-efficiency, low-cost, and multifunctional solar-driven photocatalytic systems.

    Recent advances in g-C3N4-based S-scheme photocatalyst
    Shike Wang, Man Lang, Feng Wang, Pengfei Xia, Dieqing Zhang
    2027, 92:  37-64.  DOI: 10.1016/S1872-2067(26)65252-7
    Abstract ( 250 )   HTML ( 1 )   PDF (9841KB) ( 135 )  

    Benefiting from Fermi level equilibration, interfacial band bending, and built-in electric fields, S-scheme heterostructures achieve efficient interfacial charge separation while preserving the robust redox capability of photogenerated charge carriers. This review systematically clarifies the evolutionary logic of heterojunction charge transfer models, spanning from conventional type-II and diverse Z-scheme configurations to advanced S-scheme heterostructures, and elaborates the core interfacial semiconductor physics governing S-scheme systems. A comprehensive overview is provided on the experimental characterization and theoretical verification techniques for identifying S-scheme charge transfer pathways, including density functional theory simulations, Kelvin probe force microscopy, in-situ X-ray photoelectron spectroscopy, femtosecond transient absorption spectroscopy, and electron paramagnetic resonance. On this basis, the unique superiority of graphitic carbon nitride (g-C3N4) as a reduction photocatalyst (RP) in S-scheme systems is thoroughly analyzed from the perspectives of thermodynamic band structure, interfacial charge redistribution, layered structural compatibility, and flexible surface modification. Furthermore, this paper summarizes the state-of-the-art advances of g-C3N4-based S-scheme photocatalysts for typical catalytic reactions, including photocatalytic H2 evolution, H2O2 production, photoreductive CO2 conversion, bacterial disinfection, and organic pollutant degradation. Corresponding reaction-oriented structural optimization strategies are also categorized and discussed, involving dimensional matching engineering, active site regulation, homojunction construction, and bifunctional redox coupling design. Finally, the key bottlenecks that restrict the practical application of g-C3N4-based S-scheme photocatalysts are highlighted.

    Engineering electrocatalysts for industrial-current-density alkaline hydrogen evolution: Design principles and future perspectives
    Panyong Kuang, Zhenrui Ni, Sihan Chen, Weilai Yu
    2027, 92:  65-86.  DOI: 10.1016/S1872-2067(26)65280-1
    Abstract ( 55 )   HTML ( 2 )   PDF (13997KB) ( 10 )  

    Renewable electricity-powered water electrolysis is a cornerstone technology for sustainable hydrogen (H2) production and future carbon-neutral energy systems. However, realizing economically viable large-scale H2 production requires electrocatalysts capable of sustaining high activity and long-term durability at industrial current densities, where intensified mass transport limitations, rapid gas evolution, and accelerated catalyst degradation fundamentally alter reaction kinetics and device performance. Recent advances in alkaline hydrogen evolution reaction (HER) electrocatalysts have demonstrated that efficient industrial-current-density operation relies not only on intrinsic catalytic activity but also on the synergistic optimization of catalyst architecture, interfacial mass transport, bubble dynamics, and structural stability. In this review, we provide a comprehensive overview of recent progress in alkaline HER electrocatalysts from the perspectives of morphological and interfacial engineering, intrinsic activity regulation, and durability enhancement, while establishing the relationships among catalyst structure, reaction kinetics, mass transport, gas evolution, and long-term operational stability. We further discuss challenges in bridging laboratory-scale catalyst development with practical water electrolysis, including interfacial microenvironment regulation, catalyst-layer engineering, standardized performance evaluation, and dynamic operation under renewable-energy-driven conditions. Finally, we highlight emerging design principles and future opportunities for developing highly efficient and durable HER electrocatalysts for practical green hydrogen production.

    Recent advances in metal-organic framework-based electrocatalysts for electrochemical reduction of nitrate to ammonia
    Bo Zhang, Yuye Jiao, Huijie Cheng, Jungang Hou
    2027, 92:  87-114.  DOI: 10.1016/S1872-2067(26)65269-2
    Abstract ( 54 )   HTML ( 0 )   PDF (7788KB) ( 9 )  

    Electrocatalytic conversion of nitrate (NO3-) to ammonia (NH3) represents a compelling strategy for mitigating the problem of nitrogen pollution while simultaneously enabling waste valorization, thereby offering a sustainable alternative to the energy-intensive Haber-Bosch process for NH3 synthesis. Metal-organic frameworks (MOFs), a class of porous materials composed of metal nodes and organic linkers, exhibit periodically extended network structures. Their inherent advantages, including high porosity, well-defined and uniform active sites, and tunable organic functionalities, position them as promising candidates in the field of energy and catalysis. Therefore, a systematic summary of the characteristics and recent research progress of MOF-based materials for the electroreduction reaction of NO3- holds significant importance. This review first introduces the current application of pristine MOF materials, MOF-based composites, and MOF-derived materials in the field of NO3- electroreduction reaction. It elaborates on the structural characteristics and performance differences of various types of catalysts, revealing the relationship between the catalytic activity and the composition. Subsequently, we highlight and discuss the key strategies for enhancing the performance of NO3- electroreduction reaction, thus offering theoretical insights to rationally design the catalysts with high activity and long-term stability. Furthermore, to achieve an in-depth understanding of the mechanism, this review delineates the state-of-the-art techniques employed to probe the structure of catalyst, as well as provide a reliable basis for understanding the reaction pathways of NO3- electroreduction reaction. Finally, the significance of MOF-based materials for NO3- electroreduction is summarized, and perspectives on their future development are provided.

    Communications
    Anaerobic photocatalytic dihydroxylation of propylene to propylene glycol and hydrogen at ambient temperature and pressure
    Xinyu Song, Wei Wang, Xiaomin Lao, Yitong Liu, Huanmin Liu, Peng Zhou
    2027, 92:  115-122.  DOI: 10.1016/S1872-2067(26)65228-X
    Abstract ( 76 )   HTML ( 1 )   PDF (6059KB) ( 16 )  
    Supporting Information

    The oxidant-free dihydroxylation process of olefin using water as hydroxyl source represents a highly desirable strategy for synthesizing diols as it reduces the risk of over-oxidation of the olefin and enhances the safety of the production process, however, remaining to be a challenge under mild reaction conditions. Herein, we demonstrate a photocatalytic non-oxidative dihydroxylation pathway that only can utilize sunlight, propylene, and water to directly produce 1,2-propylene glycol (1,2-PG) and hydrogen over a silver nanoparticle-loaded TiO2 (Ag-TiO2) under ambient pressure and temperature. In-situ spectroscopic analyses combining with density functional theory simulations revealed that only Ag nanoparticles can capture photogenerated holes from TiO2 instead of electrons, enabling the rate-determining hydroxylation step with a lower energy barrier (0.33 eV) at the Ag-TiO2 interface compared to other noble metal-TiO2 interfaces. A 1,2-PG yield of 177.7 μmol g-1 h-1 was achieved on Ag-TiO2 via photocatalytic non-oxidative dihydroxylation pathway under ambient conditions. In contrast, only a trace amount of 1,2-PG was detected on other noble metal-loaded catalysts. More importantly, Ag-TiO2 could consistently produce 1,2-PG (168.4 μmol g-1) and hydrogen without any decline in activity under natural sunlight irradiation. This work establishes an integrated photocatalytic route for non-oxidative dihydroxylation process of olefin coupled with hydrogen production, offering a green and scalable pathway for solar-driven olefin functionalization.

    Frontier orbital insights into the stability and activity of zero-valent single-atom catalysts
    Yuxing Xu, Yin Li, Hengwei Wang, Botao Teng, Junling Lu
    2027, 92:  123-129.  DOI: 10.1016/S1872-2067(26)65217-5
    Abstract ( 71 )   HTML ( 0 )   PDF (2999KB) ( 25 )  
    Supporting Information

    The stability of single-atom catalysts (SACs) has been a major concern for their practical applications. The orbital-level understanding of the nature of metal-support interaction is of essential importance, but has been rarely reported. Here, frontier molecular orbital (FMO) theory is employed to investigate the stabilization mechanism of a nearly zero-valent Pd1 catalyst supported on boron-doped g-C3N4 (Pd1/BCN). We demonstrate that the exceptional stability originates from enhanced frontier orbital coupling under a staggered energy-level alignment between the Pd1 atom and the support. Boron doping effectively modulates the support's band structure, enhancing both the charge transfer from the highest occupied molecular orbital of Pd1 atom to the support's lowest unoccupied state and the electron back-donation from the support's highest occupied state to the lowest unoccupied molecular orbital of Pd1. This balanced dual coupling minimizes net charge transfer, thereby stabilizing Pd1 atoms in a nearly zero-valent state. The FMO theory further consistently rationalizes the superior H2 activation kinetics and thermodynamics on electron-enriched Pd1/BCN, establishing it as a valuable framework for the rational design of SACs with optimized activity and stability.

    Articles
    Engineering local electronic asymmetry in isostructural covalent organic frameworks to maximize charge separation for H2O2 photosynthesis
    Yangpeng Zhang, You Wu, Qianfeng Gu, Yue Gao, Chencheng Cai, Haoyu Liu, Yuehan Zhao, Zhonghua Li, Feng-Ming Zhang, Yang Liu, Qichun Zhang
    2027, 92:  130-144.  DOI: 10.1016/S1872-2067(26)65194-7
    Abstract ( 166 )   HTML ( 2 )   PDF (7184KB) ( 39 )  
    Supporting Information

    Photosynthesis of H2O2 from water and air offers a sustainable alternative to anthraquinone process. Maximizing photogenerated charge separation is pivotal for high photocatalytic efficiency in covalent organic frameworks, yet a comprehensive understanding of the interplay between global framework symmetry and local electronic asymmetry remains elusive. Herein, three isostructural β-ketoenamine- linked COFs (TAB-COF, TAM-COF, and TAT-COF) were constructed from triamine linkers with distinct N-heterocyclic cores to reveal how the synergy between global framework symmetry and local electronic asymmetry governs charge separation and H2O2 photosynthesis, while yielding frameworks with precisely modulated symmetries (D3h, C2v, and D3h) and electronic properties. TAT-COF, coupling a high-symmetry D3h framework with strongly electron-deficient triazine core, delivered a remarkable rate of 6497 μmol·g-1·h-1 for H2O2 evolution in pure water, further boosted to 69290 μmol·g-1·h-1 in benzyl alcohol, substantially outperforming most reported COF photocatalysts. Time-dependent density functional theory calculations reveal that this exceptional performance originates from the co-optimization of global symmetry, which enables ordered long-range charge transport, and strong local electronic asymmetry, which generates a powerful internal electric field for instantaneous exciton dissociation. Theoretical and in-situ spectroscopic results further associate this optimized charge- separation behavior with a dual-pathway reaction mechanism and the exceptional H2O2 production activity of TAT-COF. This work establishes a concise design principle for advanced photocatalysts: balancing global symmetry with local electronic asymmetry is critical for simultaneously promoting charge separation, dual-pathway reactivity, and high photocatalytic efficiency.

    Methyl steric-hindrance-induced twisted acridine-based conjugated porous polymers for efficient 1O2-driven photocatalytic H2O2 production
    Guobang Li, Zhilin Wang, Mengkai Wang, Yanchun Liu, Cong Wang, Xianggang Zhou, Yonghui Wang, Xuzheng Cao, Huaqiao Tan
    2027, 92:  145-156.  DOI: 10.1016/S1872-2067(26)65227-8
    Abstract ( 87 )   HTML ( 0 )   PDF (3775KB) ( 21 )  
    Supporting Information

    The effective generation and utilization of singlet oxygen (1O2) offers a promising yet challenging route for photocatalytic hydrogen peroxide (H2O2) production. In this work, we describe a molecular engineering strategy regulated by steric hindrance to control excitonic behavior and oxygen activation pathways in conjugated porous polymers (CPPs). When methyl groups are introduced into the acridine linker (from acriflavine to acridine yellow G), a planar CPP-A converts into a twisted CPP-G which has an approximately 65° dihedral angle. This change in conformation drastically alters the photophysical properties. The planar structure of CPP-A facilitates exciton dissociation and results in the development of superoxide radical (O2•-) via charge transfer. Conversely, the twisted conformation of CPP-G assists in enhancing spin-orbit coupling and intersystem crossing efficiency. These lead to an accumulation of triplet excitons and dominant 1O2 generation through energy transfer. The electrophilic 1O2 possesses a lower adsorption energy barrier on the photocatalyst surface, favoring subsequent proton-coupled electron transfer reactions. Consequently, 1O2-dominated CPP-G exhibits a remarkable H2O2 production rate of 8.97 mmol g-1 h-1 in pure water without sacrificial agents, significantly outperforming the O2•--dominated CPP-A (5.13 mmol g-1 h-1). Furthermore, CPP-G demonstrates high efficiency and selectivity in 1O2-mediated organic transformations. This work demonstrates that steric hindrance engineering serves as a generalizable strategy, enabling a fundamental shift in the reactive oxygen species generation pathway, from charge transfer to energy transfer, via simple structural modifications in metal-free CPP photocatalytic systems.

    Decoupled active centers on crystalline carbon nitride nanoarrays for synergistic photocatalytic hydrogen peroxide synthesis
    Jiaqi Li, Haojie Song, Yuxin Gao, Xinming Zhang, Jingnan Li, Siyu Wang, Huiquan Gu, Laiyu Luo, Baojiang Jiang
    2027, 92:  157-169.  DOI: 10.1016/S1872-2067(26)65229-1
    Abstract ( 62 )   HTML ( 1 )   PDF (7861KB) ( 14 )  
    Supporting Information

    Photocatalytic hydrogen peroxide (H2O2) synthesis from water and oxygen represents a sustainable pathway for solar energy conversion. However, its efficiency is limited by rapid charge recombination, sluggish water oxidation kinetics, and insufficient proton supply. Herein, we report a photocatalyst featuring spatially separated redox centers, constructed by co-anchoring indium single atoms (In SAs) and cyano groups on highly crystalline carbon nitride nanoarrays. Structural and in situ spectroscopic analyses reveal that cyano groups act as efficient oxygen reduction reaction (ORR) sites. Concurrently, the In SAs act as water oxidation reaction (WOR) centers, which activate water molecules and supply protons for the ORR pathway. The optimized catalyst achieves a remarkable H2O2 production rate of 475 μmol g-1 h-1 under visible light, a 17-fold improvement over pristine carbon nitride. Theoretical calculations attribute this performance to an induced orbital distribution asymmetry, which facilitates directional carrier migration to respective active sites and concurrently lowers the energy barrier for the critical OOH* intermediate formation. This work establishes a dual-site design strategy to synergistically advance WOR and ORR, offering a promising route for efficient solar-driven H2O2 production.

    Heteroatom doping and surface engineering co-modulated electronic structure of prussian blue analogs for efficient overall water splitting
    Yuchan Liu, Wenjing Gao, Qintong Huang, Xiaoxiong Liu, Huihuan Ouyang, Rongkai Ye, Minsi Li, Jianqiang Hu
    2027, 92:  170-181.  DOI: 10.1016/S1872-2067(26)65222-9
    Abstract ( 67 )   HTML ( 0 )   PDF (6859KB) ( 15 )  
    Supporting Information

    Overall water splitting based on Prussian blue analogues (PBAs) electrocatalysts has attracted increasing attention due to its tunable structures, facile synthesis, reducing costs and promoting green energy applications. However, it is usually hindered by structural instability and limited active sites of electrocatalysts. Herein, we have constructed vanadium-doped and surface-phosphated NiFeP PBAs/Ni foam (V-NiFeP PBAs/NF) nanocubes (NCs) with excellent stability and catalytic performance by heteroatom doping and surface engineering strategy. Benefiting from optimized electronic structures and reconstructed active oxyhydroxides, the V-NiFeP PBAs/NF NCs as pre-catalysts exhibited exceptional oxygen evolution reaction (OER) activity in alkaline media (~257 mV at 100 mA·cm-2), which was ~42% lower than commercial RuO2 (~443 mV) and excellent stability (500 h at 50 mA·cm-2). Experimental and theoretical results indicated that the V-doping of the V-NiFeP PBAs/NF NCs could effectively modulate its electronic configuration by upward-shifting d-band center of Ni and Fe, thereby increasing its conductivity and active sites. The V-NiFeP PBAs/NF NCs had lower reaction energy barrier for lattice oxygen mechanism pathway than that for adsorption evolution mechanism pathway. The V-NiFeP PBAs/NF NCs had excellent hydrogen evolution (~170 mV at 20 mA·cm-2) and overall water splitting performance (~1.59 V at 10 mA·cm-2). This work provides a synthetic paradigm for in-situ growth of surface-engineered PBAs on conductive substrates and offers fundamental insights for designing high-performance OER catalysts.

    Dopant-enabled RuO2(110)-dominant architecture for boosting sustainable acidic oxygen evolution
    Piracha Sanwal, Jingjing Zhang, Rashid Mehmood, Linkun Sun, Xu Hu, Wen Dai, Gao Li
    2027, 92:  182-191.  DOI: 10.1016/S1872-2067(26)65270-9
    Abstract ( 53 )   HTML ( 1 )   PDF (5571KB) ( 4 )  
    Supporting Information

    Although Ru-based catalysts have been widely exploited in oxygen evolution reaction (OER), they still face significant challenges in simultaneously achieving high activity and long-term stability. Here, we propose a catalyst design strategy that engineers phases beyond conventional atomic ordering by constructing a RuO2(110)-dominated architecture (CeCo-RuO2). The resulting catalyst exhibits an overpotential of 198 mV@10 mA cm-2 and exceptional durability, operating stably for over 500 h at a current density of 100 mA cm-2 with a minimal current decay rate of 0.04 mV h-1. The engineered RuO2 phase architecture and atomic arrangement induce electronic structure reconfiguration, thereby optimizing active-site density and structural robustness. In-situ characterization combined with density functional theory calculations reveal that CeCo-RuO2 preferentially follows adsorbate evolution mechanism with reduced energy barriers, thereby suppressing lattice oxygen participation and mitigating structural instability. In all, we demonstrate dopant-enabled phase confinement as an effective design strategy for developing active and durable OER catalysts.

    Oxygen-bridged Ni-O-Co electronic conduits enable spatially partitioned dual active sites for ampere-scale hydrazine electrolysis
    Qiangqiang Sun, Zhao Li, Meina Jiang, Dan Wang, Baoyue Cao, Ning Nan, Chun Li, Chunsheng Zhou, Chuan Zhao, Yibing Li
    2027, 92:  192-205.  DOI: 10.1016/S1872-2067(26)65268-0
    Abstract ( 96 )   HTML ( 0 )   PDF (14246KB) ( 22 )  
    Supporting Information

    Hydrazine electrolysis offers a sustainable route for energy-efficient hydrogen production, yet the development of electrocatalysts capable of ampere-scale performance remains challenging. Herein, we show a Janus tubular Co3O4/NiCu heterostructure supported on nickel foam (Co3O4/NiCu/NF) via a voltammetric-assisted alloying-selective dealloying process. The generated oxygen-bridged Ni-O-Co dual active sites at the heterointerface act as a bifunctional catalyst for the hydrazine oxidation reaction (HzOR) and hydrogen evolution reaction (HER) at ultralow overpotentials of 451 mV and 263 mV to attain 3 A cm-2, respectively. Only 2.249 V cell voltage is required for overall hydrazine splitting (OHzS) to achieve the same industrial-scale current density alongside excellent stability. Synchrotron X-ray absorption spectroscopy and density functional theory calculations reveal that the Ni-O-Co bridges spatially anchor Ni and Co as adjacent dual sites and induce interfacial charge redistribution. Oxygen functions as an electronic conduit to synergistically modulate the catalytic functions: Co sites receive electrons via the unique Ni-O-Co sites to optimize H adsorption for HER, whilst Ni sites with higher oxidized states facilitate N2H4 dehydrogenation for HzOR. This work offers a robust strategy for designing dual-site heterostructured electrocatalysts for ampere-level hydrogen production.

    Asymmetric nickel single-atom and nitrogen vacancy dual sites on g-C3N4 for efficient CO2 photoreduction to CO
    Lijun Wu, Yu Xia, Chaogang Ban, Yang Wang, Xiaoxing Wang, Lujie Ruan, Zilong Zhang, Yaohan Du, Jiangping Ma, Li Hu, Haodi Ran, Ao Zhang, Xiaoping Tao, Liyong Gan, Xiaoyuan Zhou
    2027, 92:  206-218.  DOI: 10.1016/S1872-2067(26)65255-2
    Abstract ( 178 )   HTML ( 1 )   PDF (8505KB) ( 15 )  
    Supporting Information

    Photocatalytic CO2 reduction using pure water offers a sustainable route for solar‐to‐fuel conversion, yet its efficiency is limited by sluggish charge separation and the insufficient activation of CO2 and H2O. Here, we construct asymmetric dual sites on graphitic carbon nitride by coupling nickel single atoms with nitrogen vacancies (Ni1-NV/CN). The optimized catalyst achieves an outstanding CO production rate of 55.32 μmol g-1 h-1 without sacrificial agents, representing an 8.3-fold enhancement over the nitrogen-deficient graphitic carbon nitride (NV/CN) with excellent CO selectivity. Mechanistic investigations reveal a clear functional asymmetry: Ni single atoms serve as high-affinity centers for CO2 adsorption and activation, whereas nitrogen vacancies promote H2O adsorption and proton generation. This spatial cooperation accelerates proton-coupled electron-transfer steps, enhances charge separation, and substantially lowers the free-energy barrier for *COOH formation. This work establishes asymmetric dual-site engineering as an effective strategy for activating multi-molecule substrates in photocatalytic CO2 conversion and offers mechanistic insights for designing advanced metal-defect architectures.

    WO3/PHI S-scheme heterojunction with photoinduced dual defects for highly selective photocatalytic CO2-to-CO conversion
    Mengxuan Liu, Rongxin Zhang, Xin Zhang, Yanfeng Zhang
    2027, 92:  219-236.  DOI: 10.1016/S1872-2067(26)65232-1
    Abstract ( 109 )   HTML ( 1 )   PDF (12037KB) ( 27 )  
    Supporting Information

    Photocatalytic CO2 reduction, which converts solar energy into chemical energy, represents a highly promising strategy. However, conversion efficiency is often constrained by the poor adsorption and activation of CO2 and H2O molecules. Here, we report an oxygen and nitrogen-vacancy-enriched WO3/poly(heptazine imide) (PHI) S-scheme heterojunction for efficient and selective CO2 photoreduction. The optimized WO3/PHI heterojunction delivers a CO evolution rate of 129.2 μmol g-1 h-1 with 96.1% selectivity, corresponding to 7.9- and 6.5-fold enhancements over pristine WO3 and PHI, respectively. Experiments and theoretical calculations demonstrate that the in-situ formed and photoinduced dual vacancies facilitate adsorption of CO2 and H2O, thereby improving the efficiency of photocatalytic CO2 reduction. The interfacial W-N bonds serve as critical channels to accelerate interfacial charge transfer across the WO3/PHI heterojunction. The S-scheme charge-transfer pathway, in concert with a dual-defect interfacial architecture, promotes charge transport, substantially boosting CO2 reduction performance. This work provides mechanistic insight into defect-assisted S-scheme photocatalysis and offers a rational strategy for designing efficient photocatalysts for solar fuel production.

    Plasmonic hotspots-mediated C-C coupling drives selective CO2-to-C2H6 photoreduction
    Xiaolei Liu, Yinlan Liang, Jinyu Li, Huiliang Li, Zhaoke Zheng, Zeyan Wang, Jianmin Luo, Juan Li, Baojiang Jiang, Baojun Li, Zaizhu Lou
    2027, 92:  237-247.  DOI: 10.1016/S1872-2067(26)65262-X
    Abstract ( 61 )   HTML ( 1 )   PDF (9576KB) ( 8 )  
    Supporting Information

    Photocatalytic CO2 reduction to C2+ products remains challenging due to inefficient C-C coupling. Herein, we design a plasmonic heterostructure comprising Au nanoparticles (NPs) and Ni-doped W18O49 (Ni-W18O49) nanowires for selective CO2-to-C2H6 photoreduction with H2O. Plasmonic hotspots generated at the Au/Ni-W18O49 interface concentrate optical energy via localized electromagnetic field enhancement, greatly boosting hot-electron generation and enrichment at the active sites. Meanwhile, Ni dopants in W18O49 promote the formation of *CHO intermediates, while adjacent Au NPs favor the generation and stabilization of *CO species. Crucially, these plasmonic hotspots mediate the synergy between the dual active sites, driving efficient C-C coupling by forming the critical *COCHO intermediate and enabling selective C2H6 formation. Therefore, the optimized Au1.2/Ni0.32-W18O49 catalyst attains a C2H6 yield of 25.41 μmol g-1 h-1 with 72.13% selectivity, a performance unattainable with bare Ni-W18O49. This work establishes engineered plasmonic hotspots as a potent strategy to advance CO2-to-C2+ photosynthesis.

    Cobalt-coordinated covalent organic framework as a versatile water oxidation catalyst in dye-sensitized photoelectrochemical cells for solar water splitting
    Zizhan Liang, Xiaolin Wang, Zhilin Hong, Xinao Li, Xinbang Peng, Lilin Tan, Limin Xiao, Yecheng Zhou, Junmin Liu
    2027, 92:  248-261.  DOI: 10.1016/S1872-2067(26)65196-0
    Abstract ( 106 )   HTML ( 0 )   PDF (9144KB) ( 21 )  
    Supporting Information

    The development of efficient water oxidation catalysts is crucial for advancing dye-sensitized photoelectrochemical cells toward practical solar-driven water splitting. Herein, we synthesize a cobalt-coordinated covalent organic framework, TFPP-Co-COF, and integrate it with photosensitizers to construct high-performance photoanodes. The optimized electrode achieves an initial photocurrent density of 800 μA/cm2 at 0 V vs. Ag/AgCl, maintaining 700 μA/cm2 after 200 s with a Faradaic efficiency of 88.8%. Even under unbiased conditions (0 V vs. NHE), it delivers 450 μA/cm2 (71.6% FE) and retains 400 μA/cm2 over 25 min. Mechanistic studies confirm efficient electron transfer from TFPP-Co-COF to the photosensitizer. Theoretical calculations further reveal that water oxidation proceeds through a water nucleophilic attack pathway at cobalt sites, exhibiting a lower energy barrier than metal-free alternatives. The superior performance over amorphous analogs originates from enhanced crystallinity, high surface area, and abundant cobalt active sites that collectively promote charge separation and catalytic activity. TFPP-Co-COF also demonstrates excellent compatibility with pure organic dyes C4BTP and DT1, highlighting its versatility. This work pioneers COF application in DSPECs, providing a promising pathway for efficient solar fuel production.

    Tailoring the d-band center of CoSe2 cocatalyst via Ni-doping-induced p-d hybridization perturbation for enhanced photocatalytic H2 production
    Shiwen Du, Quanmei Li, Na Lu, Wei Lu, Jindou Huang, Zhenyi Zhang
    2027, 92:  262-273.  DOI: 10.1016/S1872-2067(26)65236-9
    Abstract ( 80 )   HTML ( 0 )   PDF (4184KB) ( 29 )  
    Supporting Information

    Loading cocatalysts plays a crucial role in enhancing the photocatalytic H2 evolution performance of semiconductors. However, due to the undesirable electronic configuration of active sites, the H2 evolution activity of intrinsic cocatalysts remains significantly limited. Herein, we propose a novel strategy for modulating the d-band center (εd) of transition metal active sites by transition metal heteroatom doping. The introduced heteroatoms perturb the nonmetal p-band configuration in the cocatalyst and optimize the orbital hybridization between the nonmetal p-band and the d-band of active sites. Such regulation enables precise modulation of the εd of active sites and the H* adsorption-desorption equilibrium, thereby ultimately boosting the photocatalytic H2 evolution performance. This tactic is demonstrated by the Ni-doped CoSe2 (NCS) cocatalyst and Bi4Ti3O12 (BTO) semiconductor. Specifically, the introduction of Ni with higher electronegativity effectively promotes the rearrangement of outer-shell electrons of Co active sites, leading to an upward shift of its εd and thereby enhancing the H* adsorption capacity. Accordingly, the sample of BTO loaded with NCS delivers excellent photocatalytic H2-evolution activity, which is approximately 20.1 and 1.8 times higher than that of pristine BTO and CoSe2/BTO samples, respectively. This work provides a new insight into the rational design of high-efficiency H2 evolution cocatalysts.

    Regulating carrier dynamics in CdS via controlled sulfur vacancy engineering: Insights from femtosecond transient absorption spectroscopy
    Zheng Meng, Chenbin Ai, Jianjun Zhang, Bowen He, Liuyang Zhang, Zhen Wu, Serguei Filatov, Dmitry Selishchev, Jahan B. Ghasemi, Jiaguo Yu
    2027, 92:  274-284.  DOI: 10.1016/S1872-2067(26)65278-3
    Abstract ( 50 )   HTML ( 1 )   PDF (12342KB) ( 14 )  
    Supporting Information

    Severe electron-hole recombination remains a fundamental bottleneck limiting the efficiency of semiconductor photocatalysts. Engineering defect states, particularly sulfur vacancies (Sv), offers an effective strategy to tailor local electronic structures and regulate charge carrier dynamics. Herein, CdS hollow spheres with tunable Sv concentrations were constructed through thermal treatment, yielding a non-monotonic (volcano-type) dependence of defect density. Ultrafast femtosecond transient absorption spectroscopy reveals that a moderate Sv concentration introduces shallow electron-trapping states, thereby prolonging the lifetime of photogenerated electrons (up to 1112.0 ps in acetonitrile) and suppressing recombination. In contrast, excessive Sv accumulation generates recombination centers, resulting in accelerated carrier decay. Kelvin probe force microscopy further demonstrates that the optimally engineered sample exhibits the lowest surface photovoltage (77 mV), indicative of efficient electron accumulation at surface active sites. More importantly, this work identifies a concentration-dependent dual role of sulfur vacancies—evolving from charge-separation promoters to recombination centers—and elucidates the interplay between defect-mediated carrier trapping and surface reaction kinetics in aqueous environments. These findings establish a refined mechanistic framework for defect engineering and provide strategic guidance for the rational design of high-performance CdS-based photocatalysts.

    Zn vacancy-anchored Ag clusters on single-unit-cell ZnIn2S4 nanosheets for efficient dual-function photocatalysis
    Lifang Yang, Yuge Zhang, Yong Liu, Zhijun Yang, Guanghui Li, Jianwei Zhou, Sugang Meng, Raf Dewil
    2027, 92:  285-299.  DOI: 10.1016/S1872-2067(26)65246-1
    Abstract ( 101 )   HTML ( 2 )   PDF (15559KB) ( 22 )  
    Supporting Information

    Dual-function photocatalysis coupling reduction reactions with organic selective oxidation offers a sustainable strategy for energy and environmental challenges. Herein, we demonstrate a rational design of ultrathin, one-unit-cell Ag/ZnIn2S4-VZn (Ag/ZIS-VZn) nanosheets, featuring zinc vacancies (VZn) as high-affinity anchoring sites for Ag clusters. This unique configuration promotes ohmic contact between Ag clusters and the VZn-rich ZnIn2S4 substrate. Combined experiments, in situ characterizations, and theoretical studies reveal that this Ag-VZn coupling fundamentally reconfigures the charge-carrier dynamics, ensuring efficient spatial decoupling and establishing an optimized landscape for surface catalysis. Consequently, the Ag/ZIS-VZn architecture exhibits exceptional bifunctional performance for the selective oxidative valorization of phenylcarbinol into value-added chemicals, concurrently coupled with efficient hydrogen evolution, outperforming the reference benchmarks of ZIS, Ag/ZIS and ZIS-VZn. Remarkably, the integration of CO2 into the photocatalytic system enables a tandem process, coupling phenylcarbinol oxidation with CO2 photoreduction. This synergy is underpinned by strengthened CO2 adsorption and a lowered energy barrier for the rate-limiting formation of *COOH intermediate. Furthermore, 1%Ag/ZIS-VZn exhibits outstanding durability, retaining performance over 18 cycles. By elucidating the cooperative mechanism between metal vacancies and anchored clusters, this study establishes a design blueprint for high-performance, durable photocatalysts capable of steering complex, multi-component solar-driven transformations.

    Enhanced CO2 hydrogenation to heavy hydrocarbons through diffusion-limited intermediate enrichment: Semi-open vs. open systems
    Lisheng Guo, Liru Zheng, Xinyu Zhang, Xin Wang, Jie Cheng, Mengdie Cai, Na Liu, Kaige Wang, Youfeng Tian, Jie Li, Song Sun
    2027, 92:  300-315.  DOI: 10.1016/S1872-2067(26)65161-3
    Abstract ( 68 )   HTML ( 0 )   PDF (8351KB) ( 18 )  
    Supporting Information

    Precisely tuning the nanoconfined microenvironment to optimize heterogeneous catalytic reaction pathways represents one of the critical challenges for achieving efficient and directional CO2 conversion. In this study, we systematically investigated the impact of confinement dimensions on CO2 hydrogenation performance by constructing a series of semi-open porous carbon spheres (MPCSx) with tunable pore sizes (x = 0, 10, 14, 30, 100 nm) supported CoFe alloy catalysts. Experimental results revealed that the catalyst Fe10Co2@MPCS14 with a pore size of 14 nm exhibits C5+ selectivity was twice that of open systems. To gain deeper insights into the structural origins of this optimal performance, we further elucidated the underlying microscopic mechanisms by combining Finite element method simulation with in-situ diffuse reflectance Fourier transform infrared spectroscopy. The 14 nm semi-open structure established a unique “enrichment-diffusion” dynamic equilibrium around the active sites, which not only substantially enhanced the local concentrations of CO2/H2 but also effectively stabilized key C1 intermediates such as HCOO*, HCO3*, and CO3*. thereby promoting C-C coupling and carbon chain growth. This work provides important mechanistic insights into the regulation of selectivity in heterogeneous catalytic reactions through spatial confinement microenvironment modulation.

    Precise tailoring of Cu-ZrO2 interfaces for CO2 hydrogenation to methanol
    Jie Zhao, Guangyuan Xu, Si Chen, Huan Yan
    2027, 92:  316-327.  DOI: 10.1016/S1872-2067(26)65168-6
    Abstract ( 134 )   HTML ( 0 )   PDF (12365KB) ( 32 )  
    Supporting Information

    The rational design of metal-oxide interfaces is crucial for efficient CO2 hydrogenation to methanol. Here, highly dispersed ZrO2 coating was precisely modulated on the Cu/Al2O3 via atomic layer deposition, obtaining a uniform and well-defined Cu-ZrO2 interfaces. Compared with traditional methods, the optimized Cu/Al2O3@2ZrO2-A catalyst exhibited significantly enhanced CO2 conversion and methanol selectivity, achieving a methanol space-time yield of 0.537 gMeOH g-cat-1 h-1 and 9.59 g-MeOH gCu-1 h-1 at 240 °C, along with long-term stability over 400 h. Structural and electronic characterizations revealed that uniform Cu-ZrO2 interfaces with highly dispersed and vacancy-rich ZrO2 coating were achieved on the Cu/Al2O3@2ZrO2-A catalyst, demonstrating high CO2 adsorption and activation. In situ spectroscopic studies demonstrated that the uniform and robust Cu-ZrO2 interface with oxygen vacancy-rich ZrO2 coating could enhance the formation and transformation of formate intermediate. This work provides a potentially efficient strategy for precisely modulating the interface for other challenging reactions.

    Nano-confinement and hydrophobic ionic-liquid engineering for methanol-selective CO2 electroreduction from dilute streams
    Zhen Zhang, Xiangya Zhang, Mai Zhang, Yifei Yang, Yitong Wu, Linlin Zhang, Jianjun Liao, Chunfeng Song
    2027, 92:  328-339.  DOI: 10.1016/S1872-2067(26)65253-9
    Abstract ( 96 )   HTML ( 1 )   PDF (3846KB) ( 17 )  
    Supporting Information

    The electrocatalytic reduction of dilute carbon dioxide (CO2) to methanol faces dual challenges of sluggish reaction kinetics and the competing hydrogen evolution reaction (HER). This study presents a nanoconfined ionic liquid catalyst strategy, in which Cu2‒xSe nanocrystals are anchored as catalytic engines within mesoporous nitrogen-doped carbon nanofibers (CSNC), followed by impregnation with the ionic liquid [Emim][BF4] to modulate the reaction microenvironment. The multilevel pore structure of the catalyst enables local enrichment of CO2 through a nanoconfinement effect, while the homogeneous distribution of the ionic liquid combined with a hydrophobic interface collectively suppresses HER, thereby enhancing methanol selectivity. In an H-type cell with 0.5 mol/L KHCO3, CSNC/[Emim][BF4] achieves a peak methanol Faradaic efficiency (FE) of 79.4% at -0.52 V vs. RHE, sustaining > 70% FE over 48 h. In a flow cell with 1 mol/L KOH, FE reaches 78.8% under pure CO2 and 72.3% under 10% CO2 (simulating flue gas), 1.67 times higher than CSNC alone, with stable performance over 24 h. In situ spectroscopy and feedstock-switching experiments reveal N-coordination stabilizes *COOH intermediates, favoring methanol via the formate pathway. This strategy advances selective, stable CO2-to-methanol conversion from dilute streams, promoting sustainable carbon utilization and industrial flue gas valorization.

    Poly(ionic liquid)-directed pyrrolic-N-rich carbon for trace-Pd anchoring toward self-optimizing CO2-to-DMC electrocatalysis
    Yong Li, Yifan Liu, Cui Qiao, Mengqian Fu, Zifeng Yang, Yunong Li, Suojiang Zhang
    2027, 92:  340-354.  DOI: 10.1016/S1872-2067(26)65267-9
    Abstract ( 99 )   HTML ( 0 )   PDF (6606KB) ( 23 )  
    Supporting Information

    The electrocatalytic conversion of CO2 and methanol to dimethyl carbonate (DMC) offers a sustainable route for CO2 utilization, circumventing the thermodynamic constraints of thermal processes. However, current redox-neutral electrocatalytic systems face challenges in achieving catalyst-electrolyte compatibility and precise active-site control under dynamic electrochemical conditions. Here we report a self-optimizing electrocatalytic system for CO2 to DMC conversion, enabled by a pyrrolic-N-rich carbon cathode derived from bromide-containing poly(ionic liquids) (PILs). Trace Pd ions deliberately introduced into the electrolyte are in-situ captured by pyrrolic nitrogen motifs, achieving the dynamic catalyst evolution with reaction progression. The pyrrolic nitrogen dynamically modulates the electronic structure of anchored Pd species, refining active-site dispersion and synergistically coupling CO2 electroreduction with methanol carbonylation to yield DMC selectively. The system achieves a Faradaic efficiency of 75% for DMC over 6 h with minimal Pd usage, outperforming conventional pre-fabricated catalysts. This work unveils a "design-for-dynamics" paradigm wherein PIL-directed precision synthesis encodes dynamic responsiveness into the catalytic interface for sustainable CO2 conversion.

    Uniform cobalt sub-nanometer clusters catalysts for nitrate-to-ammonia electrosynthesis under industrial conditions
    Shangchun Li, Beibei Xu, Jingyu Lu, Shijie Zhao, Qingqing Zhang, Rengui Li, Jing Zhang
    2027, 92:  355-364.  DOI: 10.1016/S1872-2067(26)65199-6
    Abstract ( 77 )   HTML ( 1 )   PDF (6713KB) ( 24 )  
    Supporting Information

    The practical application of electrocatalytic nitrate reduction to ammonia (NO3RR) is limited by insufficient selectivity and stability under industrial current densities, primarily due to competitive side reactions and catalyst structural degradation. A critical challenge lies in the rational design of stable and active sub-nanometer metal clusters, where achieving strong yet precise anchoring of metal atoms to specific nitrogen sites within a carbon matrix is essential to prevent aggregation and tune the electronic structure for optimal performance. In this work, by precisely engineering the local coordination environment of nitrogen dopants, we created tailored anchoring sites that effectively stabilize cobalt atoms, thereby fostering the formation of well-defined cobalt sub-nanometer clusters (Co SNC). Mechanistic investigations reveal that the optimized Co SNC sites facilitate the selective hydrogenation of the key *NH2OH intermediate to NH3 via adsorbed hydrogen (*Hads). Consequently, the engineered catalyst achieves an exceptional ammonia production rate of 6.9 mol h-1 gcat-1 with a Faradaic efficiency of 91.4% at -0.6 V vs. RHE. Moreover, it demonstrates outstanding durability, operating steadily for over 100 h at an industrial-level current density of 500 mA cm-2@2.09 V in a membrane electrode assembly. This work elucidates the critical role of enhancing the dispersion of metal active sites, providing a viable strategy for designing efficient and stable electrocatalysts toward nitrate-to-ammonia conversion and related processes.

    Co(OH)2/Cu NWs with interface modification of hypophosphite ions for sustainable ammonia synthesis from nitrate at ~2.8 A·cm-2
    Ruihan Zhou, Tixuan Xia, Yingchen Yang, Dongbo Xu, Weidong Shi
    2027, 92:  365-375.  DOI: 10.1016/S1872-2067(26)65241-2
    Abstract ( 47 )   HTML ( 0 )   PDF (4704KB) ( 5 )  
    Supporting Information

    Ammonia production via nitrate electroreduction (NO3RR) offers an environmentally benign alternative, yet it is still extremely difficult to simultaneously attain superior Faradaic efficiency and long-lasting durability when operating at the high current densities. Herein, we report a self-supporting electrode comprising Cu nanowires uniformly coated with cobalt hydroxide incorporating surface-coordinated hypophosphite ions (H2PO2-) (denoted as P-Co(OH)2/Cu NWs), directly fabricated on Cu foam. The P-Co(OH)2/Cu NWs catalyst exhibited remarkable NO3RR performance in 0.1 mol·L-1 NaNO3/1 mol·L-1 NaOH, producing NH3 at a rate of 204.3 mg·h-1·cm-2 (-1.0 V) with a peak Faradaic efficiency of 96.8% (-0.7 V), both potentials referenced to RHE. Notably, the electrode maintains stable electroactivity and NH3 selectivity over 20 consecutive test cycles (1 h per cycle) under approximately 2.8 A·cm-2. In-situ attenuated total reflection Fourier transform infrared spectroscopy reveals that the P-Co(OH)2/Cu NWs electrode promotes enhanced deoxygenation of *NO3 and deep hydrogenation of *NOx intermediates. Cyclic voltammetry test and In-situ Raman spectroscopy demonstrate that the incorporated H2PO2- induce and stabilize the in-situ formation of metallic Co0 phase under operative electrolysis conditions. Density functional theory calculations further elucidate that the metallic Co0 phase, generated in situ during NO3RR, optimizes the adsorption energetics of *NOH, thereby substantially facilitating the energy-demanding step (*NO → *NOH) relative to Co(OH)2, while concurrently strengthening *H adsorption. This work presents a viable electrode design strategy for securing efficient and stable electrocatalytic NO3--to-NH3 conversion under high-rate operation.

    Revisiting the catalytic role of supported Na2WO4 for oxidative coupling of methane reaction
    Zejing Wei, Kunkun Wei, Xutao Chen, Yang Pan, Chengyuan Liu, Liping Xiao, Yifeng Shi, Shihui Zou, Jie Fan
    2027, 92:  376-386.  DOI: 10.1016/S1872-2067(26)65212-6
    Abstract ( 101 )   HTML ( 0 )   PDF (7559KB) ( 26 )  
    Supporting Information

    Na2WO4-based catalysts have shown great promise in the oxidative coupling of methane (OCM). Nevertheless, conventional studies have generally limited Na2WO4 loading to relatively low levels (<10 wt%), thereby revealing only the role of the Na-WOₓ species in activating methane to generate methyl radicals. Herein, we systematically investigate the loading-dependent structural evolution of Na2WO4 over an exceptionally wide range (0-50 wt%) and reveal a dual role for Na2WO4, i.e., surface Na-WOx sites for CH4 activation and small Na2WO4 clusters for heterogeneous •CH3 coupling. This finding revises the traditional heterogeneous-homogeneous mechanism for OCM and fills the knowledge gap regarding the structure-dependent catalytic function of Na2WO4 across a broad loading range. The in-depth understanding of these structure-dependent catalytic functions of various Na2WO4 species will substantially contribute to the design of more efficient catalysts to break through the inherent C2 yield limitation of conventional OCM processes.

    Mechanistic switch in methane combustion on PdOₓ catalysts mediated by oxygen vacancies
    Yanyan Chen, Sen Wang, Shanhui Zhu, He Wang, Mei Dong, Zhangfeng Qin, Jianguo Wang, Weibin Fan
    2027, 92:  387-396.  DOI: 10.1016/S1872-2067(26)65197-2
    Abstract ( 74 )   HTML ( 0 )   PDF (3651KB) ( 33 )  
    Supporting Information

    The superior activity of PdOx catalysts for methane combustion is intimately linked to oxygen vacancies. However, a molecular-level understanding of how these defects govern the reaction under working conditions remains elusive. Here, we develop a novel “two site-pairs” microkinetic strategy integrated with density functional theory (DFT) and ab initio molecular dynamics (AIMD) simulations to unravel their role. AIMD simulations reveal the structural evolution from Pd(100) to a PdO(100) film and finally to the PdO(101) facet. DFT calculations identify Pd atoms adjacent to oxygen vacancies (Pdₒᵥ) as the primary active centers for facile CH4 C-H bond cleavage on both PdOᵥ(101) and PdOᵥ(100) surfaces, while neighboring lattice oxygen atoms promote subsequent oxidation. This cooperative dual-site mechanism is central to the Mars-van Krevelen (MvK) pathway. Crucially, microkinetic modeling captures a temperature-driven switch in the dominant reaction mechanism: from the Langmuir-Hinshelwood mechanism prevalent on pristine PdO(101) to the Pdₒᵥ-mediated MvK mechanism that dominates on oxygen-deficient surfaces. The high activity of Pdov sites originates from their accessible frontier orbitals and the formation of covalent Pdov-C bonds in the transition states. Collectively, our work establishes a defect-mediated catalytic cycle, providing a fundamental basis for designing high-performance catalysts via oxygen vacancy engineering.

    Dopant-induced particle size control governs carbon yield in methane pyrolysis
    Daniël P. C. van Eck, Suzan E. Schoemaker, Stefan Bismeijer, Erik Betz-Güttner, Tom A. J. Welling, Petra E. de Jongh
    2027, 92:  397-409.  DOI: 10.1016/S1872-2067(26)65277-1
    Abstract ( 12 )   HTML ( 0 )   PDF (4731KB) ( 1 )  
    Supporting Information

    Methane pyrolysis is a promising route for low-emission H2 production with valuable nanostructured carbon as coproduct. Tuning catalyst performance through the addition of dopants has transformed many catalytic reactions, yet their role in methane pyrolysis remains largely unexplored. In this study, the role of three alkali oxide dopants (MgO, Na2O and K2O) is investigated in Ni-catalysed methane pyrolysis. The dopants greatly affected the lifetime and carbon yield of Ni/C in a thermogravimetric reactor: Mg-doping resulted in a threefold increase in carbon yield and more than double the catalyst lifetime, while K-doping decreased the carbon yield by around 50% at a concentration of 5 at%. This unexpectedly large effect of the dopants was elucidated with high-resolution electron microscopy. We revealed that the dopants remained well-dispersed on the support during catalysis and that they control particle growth at the start of the reaction there. Mg- and Na-doping enhanced Ni particle growth from 5‒6 nm to a more optimal size of 10‒12 nm, whereas K-doping stabilizes smaller Ni particles (7 nm). The final carbon yield was found to be critically dependent on the active Ni particle size, suggesting that the particle growth mediated by the dopants defines for a large part the overall performance of the catalyst in methane pyrolysis.

    Pt1/FeOx single-atom catalyst with dual-valent Pt sites for water-promoted hydrogenation of nitroarenes
    Anqi Li, Yusen Chen, Xiaoli Pan, Yujing Ren, Hongchen Cao, Guiyue Bi, Xiaofeng Yang, Aiqin Wang, Tao Zhang
    2027, 92:  410-419.  DOI: 10.1016/S1872-2067(26)65240-0
    Abstract ( 66 )   HTML ( 1 )   PDF (3139KB) ( 12 )  
    Supporting Information

    Single-atom catalysts (SACs) are highly attractive for selective hydrogenation owing to their excellent chemoselectivity toward specific functional group activation, yet the inherently cationic nature of isolated metal sites often limits hydrogen activation and thus compromises activity. Herein, we report a Pt1/FeOx SAC prepared via atomic layer deposition of Pt onto α-Fe2O3 nanosheets. Dual-valent Pt atoms are identified on different surface terminations of the exposed (001) facet, and theoretically demonstrated to exhibit distinct catalytic behaviors, i.e., Pt1δ+ sites enable heterolytic H2 activation whereas Pt10 sites facilitate a more kinetically favorable homolytic H2 dissociation. Moreover, a water-mediated hydrogen-bonding network substantially lowers the H2 dissociation and H* species spillover barriers and promotes remote hydrogenation. As a result, in the catalytic hydrogenation of 3-nitrostyrene, the Pt1/FeOx SAC exhibits approximately 24-fold higher activity relative to its water-free counterpart, while high selectivity toward 3-aminostyrene is maintained through preferential activation of nitro groups at adjacent low-valence Fe neighboring Pt single-atom sites. These findings reveal that the catalytic behavior of SACs strongly depends on the oxidation state of single atoms. Notably, metallic single-atom species with water-promoted hydrogen activation demonstrate superior hydrogenation efficiency, resulting in a markedly enhanced activity.

    Bidirectional catalysis of polysulfides and Li2S redox kinetics by single-atom modified S-MXene for advanced Li-S batteries
    Ke Fan, Yuqing Zhu, Yiran Ying
    2027, 92:  420-427.  DOI: 10.1016/S1872-2067(26)65250-3
    Abstract ( 78 )   HTML ( 0 )   PDF (6355KB) ( 7 )  
    Supporting Information

    The development of highly efficient catalysts for anchoring lithium polysulfides (LiPSs) and facilitating the Li2S redox process is crucial for advancing high-performance lithium-sulfur (Li-S) batteries. In this work, we introduce single-atom 3d transition metals anchored on V2NS2 MXene substrates (V2NS2-T, where T = Sc, Ti, V, Cr, Mn, Fe, Co, Ni, Cu, and Zn) as host materials to enhance the electrochemical behavior of Li-S batteries. By using density functional theory calculations, we demonstrate that all ten systems exhibit moderate adsorption strength toward LiPSs, effectively suppressing their dissolution and shuttling. Among them, V2NS2-Cr and V2NS2-Zn show the lowest Gibbs free energy barriers for the sulfur reduction reaction during discharge (0.15 and 0.17 eV, respectively). In addition, three V2NS2-T configurations exhibit impressive catalytic activity toward Li2S decomposition during charging, reducing the energy barrier from 0.66 eV (pristine V2NS2) to 0.27 eV (V2NS2-Cr), 0.27 eV (V2NS2-Fe), and 0.28 eV (V2NS2-Mn). Based on these results, the V2NS2-Cr monolayer is identified as a promising sulfur host material because of its fast charge/discharge kinetics. Moreover, we develop an energy-based ΔE descriptor to reduce the computational cost associated with determining Li2S decomposition barriers. This work provides valuable insights for designing high-potential MXene-based sulfur hosts and other two-dimensional conductive anchoring materials for Li-S batteries.

    B→Co electron donation programs Lewis-basic sites in cobalt boride for selective inner-sphere peroxymonosulfate activation
    Haoran Wei, Wei Zhao, Junlei Zhang, Guojia Yu, Fang Yang, Xiaobing Wang, Wuwan Xiong, Shijie Li
    2027, 92:  428-441.  DOI: 10.1016/S1872-2067(26)65238-2
    Abstract ( 54 )   HTML ( 0 )   PDF (8014KB) ( 5 )  
    Supporting Information

    Peroxymonosulfate (PMS)-based advanced oxidation processes are promising for contaminant removal but are often limited by nonselective radical pathways. Here we show that selective PMS activation can be achieved by regulating Lewis acid-base interactions in cobalt boride (CoxB). Electron donation from boron to cobalt modifies the electronic structure of surface Co sites, generating Lewis-basic centers that interact strongly with electrophilic oxygen atoms in PMS. This interaction promotes inner-sphere electron transfer and favors selective 1O2 generation while suppressing radical-mediated reactions. Under optimized conditions, the CoxB/PMS system removes 99.0% of oxytetracycline within 20 min, delivering a kinetic constant of 0.309 min-1 and a molar degradation rate of 14.48 min-1·mol·L-1, outperforming representative cobalt oxides. The catalyst also shows strong stability, retaining 97.6% activity after 15 cycles and sustaining > 99% removal efficiency in continuous-flow operation for 30 h (1.8 L throughput). X-ray photoelectron spectroscopy measurements reveal reversible Co valence modulation during PMS activation, whereas density functional theory calculations indicate stronger PMS adsorption and a lower potential-determining barrier on CoxB relative to Co3O4. Together, these results identify Lewis-basic Co sites as key centers for selective non-radical PMS activation and provide a strategy for designing efficient boride-based oxidation catalysts.

    Engineering imine reductases enables stereodivergent reductive amination of 4-substituted cyclohexanones
    Shunyu Zhao, Xinping Zhang, Xiangyuan Gu, Jieyu Zhou, Ye Ni
    2027, 92:  442-451.  DOI: 10.1016/S1872-2067(26)65279-5
    Abstract ( 35 )   HTML ( 2 )   PDF (3507KB) ( 6 )  
    Supporting Information

    Imine reductases (IREDs) enable sustainable routes to stereochemically defined amines. However, their synthetic application to 4-substituted cyclohexanones, important scaffolds in pharmaceutical chemistry, remains underexplored. Here, we identified a key residue in IREDs that acts as a stereochemical switch governing cis/trans selectivity, and established a feasible and efficient approach for stereodivergent synthesis of 4-substituted cyclohexylamines. Phylogeny-guided mining of IREDs homologues yielded a subset of ten enzymes capable of catalyzing reductive amination of 4-ethylcyclohexanone in high efficiency whereas distinct diastereoselectivity. MD simulations reveal that residue 127 of IRZ06 modulates substrate orientation through steric gating and hydrophobic interactions. Mutagenesis at I127 produced two complementary variants I127L and I127P. Using a range of ketones substrates and amine donors, I127L afforded cis-products up to 95% de, whereas I127P resulted trans-products with up to 92% de. Further assessment was conducted at equivalent position of residue 127 in IRZ06 across nine IREDs, demonstrating its essential role in diastereoselectivity. Specifically, residue Pro at 127 consistently induced trans-selectivity, and Leu favored cis-selectivity. This work reveals a key residue governing diastereoselectivity of IREDs, offering a generally applicable strategy for engineering IREDs to access stereochemically defined cyclohexylamine building blocks.